How to Confirm Peptide Molecular Identity in the Lab
Last Updated: October 26, 2026
Quick Summary
To confirm peptide molecular identity, calculate the expected monoisotopic mass from the stated sequence, measure the intact mass by mass spectrometry, then use tandem MS sequencing to read the residue order from fragment ions. Researchers describe how to confirm peptide molecular identity as a multi-method exercise, because intact mass alone cannot distinguish isobaric or isomeric arrangements.
- Identity, purity and content are separate measurements, and evidence for one does not substitute for another.
- Electrospray ionization produces multiply charged ions, so m/z values must be deconvolved to a neutral mass first.
- Sequence coverage is reported as the percentage of backbone bonds confirmed by at least one assigned fragment ion.
- Low-energy CID cannot distinguish leucine from isoleucine because their side-chain fragmentation is similar.
- HPLC retention time supports identity only when it is paired with mass spectrometry data.
Table of Contents
- What Peptide Identity Confirmation Actually Requires
- Peptide Mass Spectrometry: Matching Expected and Observed Mass
- LC-MS Peptide Analysis: Using Tandem MS to Confirm Sequence
- HPLC Peptide Purity: What a Chromatographic Profile Can and Cannot Prove
- Reading a Peptide Certificate of Analysis Critically
- A Step-by-Step Workflow for Identity Confirmation
- Common Analytical Pitfalls and How Orthogonal Testing Resolves Them
- Sourcing Compounds With Documented Identity Testing
- Frequently Asked Questions
What Peptide Identity Confirmation Actually Requires
Confirming peptide molecular identity means proving that a sample is the specific amino-acid sequence it claims to be, not merely that it is pure or present in the expected amount. A 2025 review by Elsayed et al. describes physical and chemical characterization of peptides and proteins as crucial for ensuring identity and purity, https://pmc.ncbi.nlm.nih.gov/articles/PMC11806371/“>regulatory guidelines for peptide and protein analysis, yet the same literature notes that no single assay covers every attribute.
That gap is where most laboratory confusion begins.
Identity Versus Purity Versus Content
These three terms describe different measurements and are frequently conflated in laboratory reports.
- Identity asks whether the molecular structure matches the target peptide sequence.
- Purity asks what fraction of the sample is the target compound versus related impurities.
- Content asks how much peptide material is present by mass or concentration.
A sample can be pure but wrong, or correct but impure. Identity testing and purity testing answer separate questions, and analytical evidence for one does not substitute for the other.
Peptide Mass Spectrometry: Matching Expected and Observed Mass
Peptide mass spectrometry is investigated as a means of confirming molecular identity by measuring the mass-to-charge ratio (m/z) of intact ions and deriving the neutral molecular mass for comparison against a calculated value. The literature describes intact-mass measurement as necessary but not sufficient for identity, a matching mass supports the molecular formula, not the sequence order. Researchers describe the comparison as a two-part exercise: first establish what mass the stated structure should produce, then explain every observed ion in the spectrum as that structure, an adduct, or an artifact.
Calculating Expected Mass From the Sequence
Expected monoisotopic mass is calculated by summing residue masses for the stated sequence, adding the mass of a water molecule to account for the free termini of an unmodified linear peptide, and then applying each declared modification as a discrete mass delta.
Representative residue and modification deltas that practitioners routinely reconcile against an observed mass include:
- Glycine, the smallest residue, contributes approximately 57 Da; tryptophan, the largest, approximately 186 Da.
- N-terminal acetylation adds approximately 42 Da; C-terminal amidation subtracts approximately 1 Da relative to the free acid.
- Oxidation of methionine adds approximately 16 Da; deamidation of asparagine or glutamine adds approximately 1 Da.
These values are widely tabulated in proteomics methods literature; the practical discipline is to enumerate every declared and plausible undeclared modification before concluding that a mass difference indicates a wrong sequence.
Charge States, Adducts, and the Isotope Envelope
Electrospray ionization produces a distribution of multiply charged ions, so the observed m/z values are not the molecular mass (Electrospray Ionization Mass Spectrometry). For a peptide of neutral mass M carrying z protons, the observed m/z is approximately (M + z)/z; the spacing between adjacent charge states in the envelope is approximately 1/z in m/z units, which allows the charge state to be inferred directly from the spectrum.
Common adducts and their approximate mass offsets, sodium (~22 Da), potassium (~38 Da), and ammonium (~17 Da), appear as satellite peaks and are investigated as expected features rather than contaminants.
A defensible mass comparison states the ionization mode, the charge states assigned, the adducts accounted for, and whether monoisotopic or average mass was used. Without those four details, a reported mass error cannot be interpreted.
Reading a Mass Shift Before Sequencing
A mass difference is investigated as a diagnostic signal rather than a verdict. The magnitude and direction of the shift narrow the fault: a difference near a single residue mass suggests a deletion or insertion; a ~16 Da offset suggests oxidation; a ~1 Da offset suggests deamidation or an amidation state error; a ~2 Da offset suggests a disulfide state error; and a shift consistent with a protecting group suggests incomplete deprotection.
LC-MS Peptide Analysis: Using Tandem MS to Confirm Sequence
LC-MS peptide analysis couples liquid chromatography to mass spectrometry, and tandem mass spectrometry (MS/MS) extends the measurement by isolating a precursor ion, fragmenting it, and recording the product-ion spectrum. The literature describes MS/MS as the technique that converts a mass match into a sequence claim, because the fragment pattern encodes the order of residues rather than merely their sum. Researchers describe sequence confirmation as an inference from which backbone bonds broke and which did not, a probabilistic reading, not a direct observation of the sequence.
Fragment Ion Nomenclature and What the Series Reveal
The convention in peptide fragmentation literature assigns b-ions and a-ions to fragments retaining the N-terminus and y-ions and x-ions to fragments retaining the C-terminus, with the subscript denoting the number of residues in the fragment. Collision-induced dissociation predominantly produces b- and y-ions, and the mass difference between consecutive ions in a series corresponds to the residue mass at that position.
Sequence Coverage, Gaps, and Isobaric Ambiguity
Sequence coverage is the fraction of backbone bonds confirmed by at least one assigned fragment ion, and it is reported as a percentage of the peptide length. Gaps in coverage, regions where no fragment ion brackets a residue, are investigated as the weak points of a sequence claim, and the literature treats a high-coverage spectrum with a single unconfirmed residue differently from a low-coverage spectrum with many.
Other interpretation issues the literature examines include:
- Incomplete fragmentation, where the precursor survives or fragments only at labile bonds, leaving the interior sequence unconfirmed.
- Internal fragments and immonium ions, which carry residue-composition information but not positional information.
- Modification localization, where a mass delta must be placed on a specific residue rather than merely detected.
- N- and C-terminal modifications, which shift entire ion series and can be mistaken for sequence errors if the specification is not applied to the expected fragments.
A matching intact mass plus a partial fragment ladder is not sequence identity. The defensible conclusion states which residues were confirmed, which were not, and why the unconfirmed positions are acceptable for the intended use.
Combining MS/MS With Orthogonal Evidence
Researchers describe MS/MS as one leg of an orthogonal confirmation strategy rather than a standalone proof. Retention-time agreement with an authenticated reference standard, intact-mass agreement, and fragment-ion agreement are investigated as independent lines of evidence, and the literature notes that agreement across methods that fail in different ways is stronger than agreement within a single method. For a broader primer on how these datasets fit together, see how to verify peptide purity.
HPLC Peptide Purity: What a Chromatographic Profile Can and Cannot Prove
HPLC peptide purity is assessed from a chromatographic profile, where the target peptide appears at a characteristic retention time and impurities appear as separate peaks. The profile quantifies purity but cannot prove identity on its own.
What chromatography can show:
- Relative purity by peak area
- Presence of related compounds and impurities
- Retention time consistency against a reference standard
What it cannot show:
- The amino-acid sequence
- Whether the main peak is the correct molecule
Retention time matching supports identity only when paired with mass data.
Reading a Peptide Certificate of Analysis Critically
A peptide Certificate of Analysis should be read as a set of separate claims, each tied to a specific analytical method. The document is only as strong as the methods behind it.
Check for:
- Batch-specific results rather than generic specifications
- The analytical method named for each attribute
- Mass spectrometry data supporting molecular identity
- Chromatographic conditions and retention time
APEX 100 RESEARCH provides a batch-specific Certificate of Analysis with every order, confirming ?99% HPLC purity and molecular identity through HPLC and mass spectrometry testing. Batch records are available for review on the certificates of analysis page.
A Step-by-Step Workflow for Identity Confirmation
A practical workflow for how to confirm peptide molecular identity sequences the methods so each result narrows the next question.
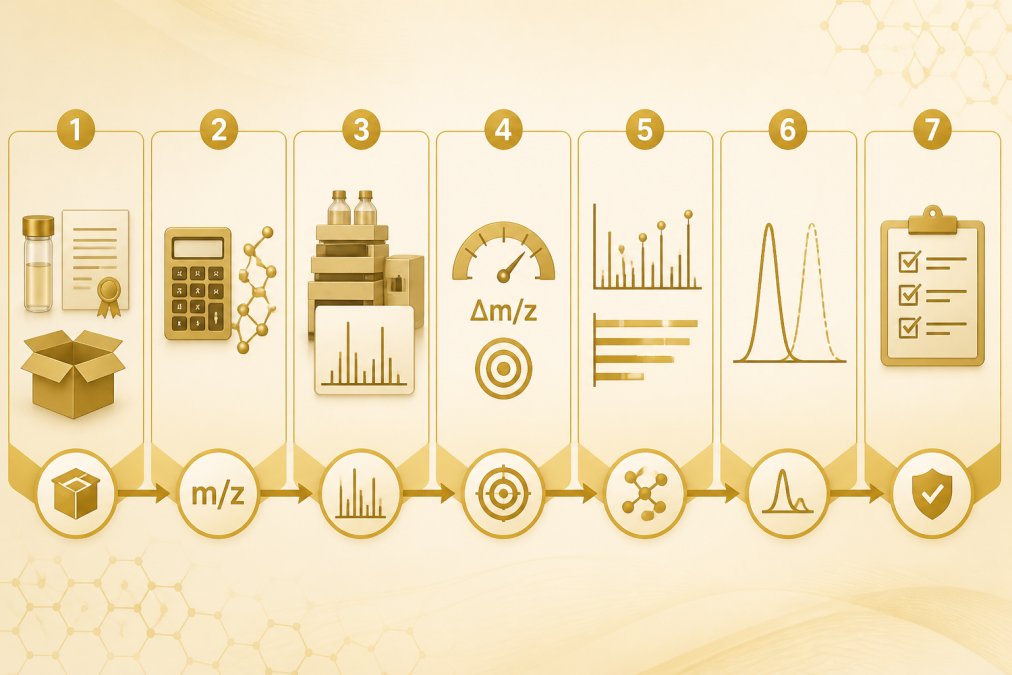
- Receive the sample and review the COA.
- Calculate expected molecular mass from the stated sequence.
- Run LC-MS to obtain observed mass.
- Compare the mass difference against method tolerance.
- Run MS/MS and assess sequence coverage from fragment ions.
- Cross-check HPLC retention time against a reference standard.
- Document the orthogonal evidence as a single record.
Quick answers on purity testing, COAs, and reconstitution logistics sit in the FAQ.
Common Analytical Pitfalls and How Orthogonal Testing Resolves Them
Most identity failures trace to method limits rather than bad samples. Orthogonal testing, which confirms a result with an independent method, resolves them.
| Pitfall | Why It Misleads | Orthogonal Fix |
|---|---|---|
| Intact mass only | Cannot resolve sequence isomers | Add MS/MS sequencing |
| Single HPLC peak | Purity is not identity | Pair with mass data |
| Tolerance too wide | Hides real mass shifts | Tighten acceptance criteria |
| No reference standard | Retention time unanchored | Run certified standard |
| One lab, one method | Method bias undetected | Independent confirmation |
A deeper treatment of third-party verification appears in the independent lab testing guide.
Sourcing Compounds With Documented Identity Testing
Sourcing decisions determine how much identity evidence arrives with the sample. Suppliers that publish batch-specific mass spectrometry and HPLC data remove the need to reconstruct provenance later. Compounds with documented testing data can be browsed in the research peptide catalog, and the Peptide Reconstitution Calculator is available for researchers working through reconstitution volumes once identity and purity documentation has been reviewed.
For research use only, not only —. Not medical advice.
Frequently Asked Questions
How can a laboratory confirm a peptide’s molecular identity?
Identity confirmation rests on orthogonal analytical evidence rather than a single result. Mass spectrometry establishes observed molecular mass, which is compared against the expected mass calculated from the declared amino-acid sequence. Tandem mass spectrometry then fragments the peptide and the resulting fragment ions are matched to the predicted sequence. HPLC provides a retention time and chromatographic profile that are compared against a reference standard. Researchers describe the combination of mass accuracy, sequence coverage, and chromatographic agreement as the basis for a defensible identity claim.
How do mass spectrometry and HPLC differ when testing peptides?
The two methods answer different questions. Mass spectrometry measures molecular mass and, in MS/MS mode, sequence-informative fragmentation, so it addresses peptide identity directly. HPLC separates components in a sample and reports a chromatographic profile with retention times, which speaks to purity and related compounds. A sample can be 99% pure by HPLC area and still be the wrong peptide entirely, so HPLC peptide purity data alone cannot confirm identity. Identity testing requires mass data, and purity testing requires chromatographic data.
Can HPLC alone confirm peptide molecular identity?
No. A chromatographic retention time is a comparative observation, not a structural measurement. It can show that a peak matches a reference standard run under identical conditions, which is useful supporting evidence, but co-elution does not prove the molecular structure. Two related compounds with similar hydrophobicity can share a retention time. Analytical practice treats HPLC as an orthogonal confirmation layer alongside mass spectrometry, not as a substitute for it.
What information should a peptide Certificate of Analysis include?
A useful peptide Certificate of Analysis identifies the target peptide and its declared sequence, states the expected molecular mass alongside the observed mass and the method used, reports HPLC purity with the chromatographic conditions and detection wavelength, and names the independent laboratory that performed the testing. Batch and lot identifiers, a testing date, and the acceptance criteria applied should also appear. Documents that report a purity percentage without stating the method or the observed mass give the reader little to verify against.
The analytical literature is clear that identity rests on orthogonal evidence, not a single assay. Laboratories that need documented molecular identity and ?99% HPLC purity, batch-specific COAs, and independently verified testing records can get started with APEX 100 RESEARCH and source compounds with traceable analytical documentation.